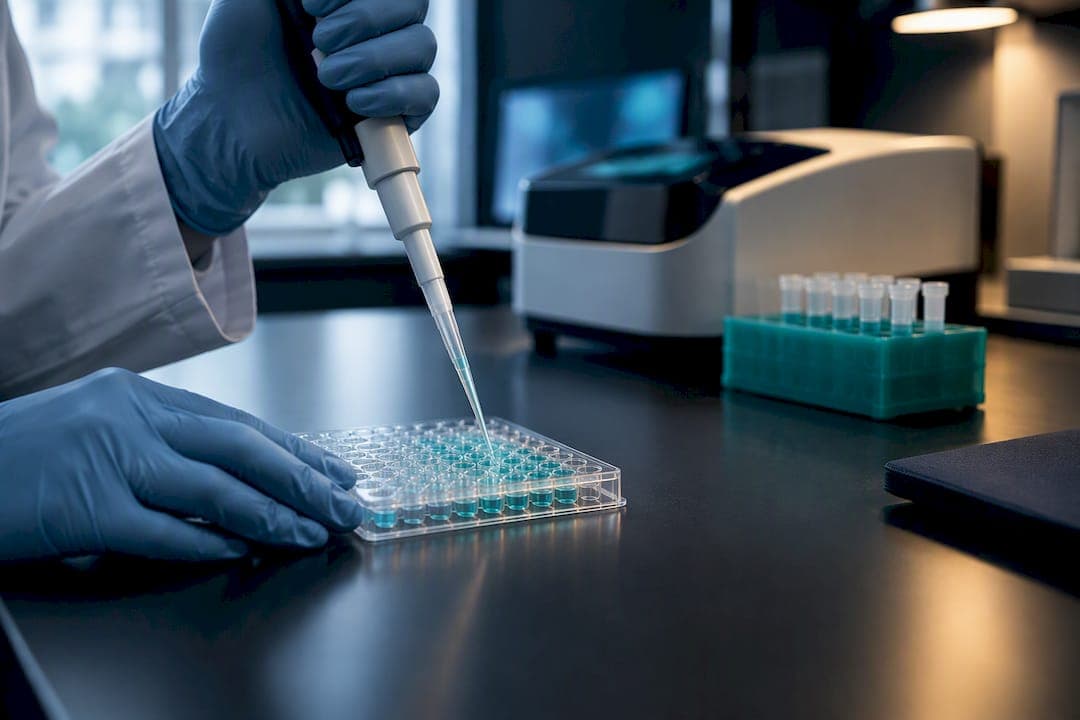
Contamination in research peptides falls into a small number of predictable categories: synthesis by-products, oxidative and structural modifications, aggregates and counter-ions, residual solvents, endotoxin, and cross-contamination from shared lab workflows. The three priorities on receipt of any lot are simple to state and easy to skip: reconcile identity against the Certificate of Analysis, validate the endotoxin method for that specific peptide matrix, and log traceability before the vial leaves the bench.
TL;DR:
- Most peptide contamination originates from synthesis impurities, oxidation, racemization, or cross-contamination, and they often escape detection by UV alone.
- Confirming identity with LC-HRMS and validating endotoxin tests in the actual peptide matrix are essential steps before use.
- Proper receipt procedures, including checking packaging, logging temperature, and assigning unique identifiers, prevent damage and mislabeling.
- Maintaining dedicated lab zones, using qualified consumables, and running procedural blanks minimize cross-contamination risks.
- Building risk-based specifications and investigating thresholds for unknown impurities or endotoxin failures improve peptide lot safety and reproducibility.
Table of Contents
- Understanding Peptide Contamination Risks in Synthetic Production
- Analytical and Endotoxin Testing: Fit-for-Purpose Approaches
- Shipment, Receipt, and Storage: Where Integrity Gets Lost
- Lab Practices That Prevent Cross-Contamination
- Setting Incoming Specifications and Investigation Triggers
- How USAPeptide Supports Contamination Risk Control
- The Real Gap Between Contamination Guidance and Lab Practice
- Verified Sourcing Starts Before the Order Ships
- Sources
- FAQ
Understanding Peptide Contamination Risks in Synthetic Production
Solid-phase peptide synthesis (SPPS) generates a predictable family of impurities, and knowing which one you're looking at narrows the investigation fast. Incomplete coupling produces deletion sequences; unwanted side reactions during deprotection leave truncated chains or protecting-group adducts still attached. These synthesis-related impurities are chemically similar to the target peptide, which is exactly why they are hard to catch.
Modification-related impurities show up after synthesis is technically complete. Oxidation, most common at methionine, tryptophan, or cysteine residues, and deamidation at asparagine or glutamine both shift mass by only a few atomic units. Racemization during coupling produces diastereomers, epimers with identical mass and near-identical polarity to the parent peptide. Structurally similar impurities like these routinely slip past HPLC-UV, because UV detection separates on hydrophobicity, not identity.
Beyond the peptide chain itself, three other contamination classes matter:
- Aggregates and oligomers, formed when hydrophobic peptides self-associate in solution or during lyophilization.
- Counter-ions, most often trifluoroacetate carried over from reverse-phase purification, which can confound downstream assays if not accounted for.
- Residual solvents, from acetonitrile to DMF, left behind when drying protocols run short.
Unrelated peptide contamination, a different sequence entirely, usually enters through shared synthesis equipment or resin cross-use. Its consequence in a bioassay is the most dangerous kind of error: an apparent activity shift or a false positive that looks like real biology until someone performs an orthogonal identity check.
Analytical and Endotoxin Testing: Fit-for-Purpose Approaches
HPLC-UV purity numbers describe how clean a peak looks, not what else is hiding under it. Because UV detection can't distinguish an epimer or a deletion sequence from the target peptide when they co-elute or sit close in retention time, a high HPLC-UV purity figure is not equivalent to a full identity or impurity assessment. LC-MS, particularly UPLC-HRMS, resolves what UV can't: it identifies isomers, epimers, and low-level related impurities by mass rather than by retention behavior alone.
A workable LC-MS workflow for incoming peptide lots looks like this:
- Run HPLC-UV first to flag any peaks worth investigating.
- Follow with LC-HRMS to confirm identity and screen for mass-matched impurities.
- Add NMR when stereochemistry or exact structural assignment is in question.
- Use SEC when aggregation is suspected, particularly for longer or hydrophobic sequences.
Response-factor differences between the target peptide and its impurities can distort quantitation, so treat LC-MS impurity percentages as directional unless calibrated against a characterized reference standard.
Endotoxin testing carries its own traps. Gel-clot, turbidimetric, and chromogenic Limulus Amebocyte Lysate assays all work on peptides, but USP General Chapter <85> and FDA guidance both require inhibition and enhancement controls run in the actual sample matrix, not a generic buffer. A peptide that inhibits the LAL reaction can mask real endotoxin; one that enhances it produces a false failure. Demonstrating method suitability, including maximum valid dilution, before trusting any endotoxin result isn't optional paperwork. It's the only way to know the number means anything.

For research work exploring immune-relevant effects, cell-based innate-immune activation assays or IIRMI methods can supplement LAL data when a peptide's own biology makes lysate results ambiguous.
Shipment, Receipt, and Storage: Where Integrity Gets Lost
The sample lifecycle between a vendor's shipping dock and your freezer is where a lot of quiet damage happens, and almost none of it shows up on a Certificate of Analysis. WHO good-practice guidance treats storage and transport conditions as something to define, monitor, and document, because an unmonitored temperature excursion is indistinguishable from a well-handled shipment until something fails downstream.
A practical receipt sequence:
- Inspect packaging condition and any temperature indicators before opening.
- Reconcile the physical lot against the COA, checking identity, quantity, and lot number.
- Log receipt time and any recorded temperature history immediately, not at end of day.
- Quarantine anything with damaged packaging or a confirmed excursion pending disposition review.
- Assign a unique accession number before the first aliquot is drawn, and enter it in the LIMS.
Aliquoting is where mislabeling risk peaks. Preserve the original vendor label alongside your accession ID, and log every transfer so a later investigation can trace a specific aliquot back to its source lot. For storage itself, validated containers, mapped freezer temperatures, and a hard rule against repeated freeze-thaw cycles matter more than most SOPs give them credit for.
Pro Tip: Build a simple retention log linking every aliquot back to its parent accession number. When a reproducibility problem shows up six months later, that log is the fastest way to rule sample handling in or out.
Guidance on validated containers and freeze-thaw limits is covered in more depth in a dedicated reference on storage temperature, and shelf-life after reconstitution gets its own stability verification protocol.
Lab Practices That Prevent Cross-Contamination
Physical layout drives a large share of contamination risk, and it's fixable without new equipment. Establishing dedicated zones for sterility-sensitive work, separate from general bench activities, reduces the chance of cross-contamination. When sharing equipment between zones is necessary, maintaining a documented clean-to-dirty workflow—moving only from lower-risk to higher-risk materials without reversing direction—helps minimize contamination risk.
Consumables deserve the same scrutiny as reagents. Dedicated pipette tips, tubes, and filters per project reduce carryover; where sharing is unavoidable, qualify the consumable lot before use rather than assuming it's inert.
Procedural blanks catch what visual inspection can't. Running a blank alongside every batch, and investigating any signal that shows up repeatedly across supposedly unrelated samples, is how one documented case traced a false-positive pattern back to a single contaminated reagent lot rather than a real biological effect. That kind of pattern rarely announces itself; it just looks like noise until someone checks.
Environmental monitoring rounds out the picture:
- Air sampling on a fixed schedule, not just after a suspected event.
- Settle plates and contact plates in sterility-relevant zones.
- Glove prints for personnel working directly with open samples.
- Establish documented alert and action limits for environmental monitoring and trend these data over time to detect gradual changes, instead of only reviewing results in isolation.
Pro Tip: Trend environmental monitoring data monthly, not just at the point of failure. A slow upward drift in contact-plate counts is often the earliest warning a zone is losing control, well before an actual failure occurs.
Setting Incoming Specifications and Investigation Triggers
A single purity percentage on a COA tells you almost nothing about whether a peptide is fit for your specific experiment. USP General Chapters <1503> and <1504> outline the fuller attribute set: identity, assay/purity, related substances, residual solvents, elemental impurities, counter-ion content, endotoxin, and aggregation. Which of these matter most depends entirely on the assay downstream.
Build specifications around risk, not habit:
- For cell-based assays sensitive to inflammatory signaling, endotoxin limits and method suitability outweigh a marginal purity difference.
- For structure-activity work, related substances and stereochemical purity matter more than residual solvent traces.
- For long-term stability studies, aggregation state and counter-ion content deserve their own acceptance criteria.
Document the rationale for any threshold that departs from a standard reference, whether tighter or looser, so a later reviewer understands the reasoning. Characterized reference standards and impurity reference materials resolve ambiguity when an unknown peak sits close to an identification threshold, something a single vendor chromatogram can't do alone.
Concrete triggers for a full investigation: an unknown impurity at or above identification threshold, a failed endotoxin suitability check, an identity mismatch on orthogonal testing, or any unaccounted gap in the storage and transport record.
How USAPeptide Supports Contamination Risk Control
A COA grading tool can help researchers check whether a supplied Certificate of Analysis supports the claims on the label, rather than taking a purity number at face value. Paired with detailed peptide molecular profiles, such tools can assist researchers in planning which orthogonal test, LC-MS, NMR, or SEC, fits a given sequence's known impurity risks.
For sourcing, USAPeptide points researchers toward ISO 17025-accredited third-party testing as a baseline for lot verification. In practice, this involves reconciling the COA against the physical lot at receipt, and using molecular profile information to decide whether additional identity confirmation is needed before use.
The Real Gap Between Contamination Guidance and Lab Practice
Most of the guidance in USP, FDA, and WHO documents isn't hard to follow. The gap is that labs treat it as reference material to consult after something goes wrong, instead of a checklist to run before a peptide is ever used. That's backward. A researcher who validates the endotoxin method for their specific peptide matrix before the first experiment, rather than trusting a vendor's generic sterility claim, has already eliminated one of the most common sources of unexplained assay variability.
The overrated step, frankly, is chasing a higher HPLC-UV purity number as a proxy for quality. Two lots at 99% purity can differ meaningfully in their impurity identity, and that difference is invisible without LC-MS or NMR behind it. The underrated step is traceability. A lot number and a receipt log sound like paperwork, but they're the only tool that lets a lab reconstruct what happened when a result doesn't reproduce six months later.
Prioritize identity confirmation and matrix-appropriate endotoxin validation before anything else. Purity is a starting point, not a conclusion.
— USAPeptide Team
Verified Sourcing Starts Before the Order Ships
USAPeptide's advantage over guessing at a vendor's claims is straightforward: its COA grading tool and peptide database let a researcher check identity and quality attributes before a lot ever reaches the bench, backed by sourcing from ISO 17025-accredited testing rather than a self-reported purity figure.
If contamination risk assessment is part of your protocol, three steps make the biggest difference: reconcile every incoming COA against the physical lot, validate your endotoxin method against the specific peptide matrix you're testing, and order from suppliers who provide characterized reference standards alongside the compound itself. USAPeptide's research peptide categories cover GLP-1 and metabolic peptides, tissue repair compounds, and growth hormone peptides with documented molecular profiles attached to each listing. Start by checking a current Certificate of Analysis against USAPeptide's grading criteria before your next order ships.
Sources
- Related impurities in peptide medicines - PubMed
- Pyrogen and Endotoxins Testing: Questions and Answers | FDA
- USP materials on peptide quality attributes
- Characterization of synthetic peptide therapeutics using LC‑MS - PubMed
FAQ
What Are the Most Common Peptide Contamination Risks?
The most common risks are synthesis-related impurities (deletions, incomplete deprotection), oxidative and racemization-related modifications, aggregates, residual solvents, endotoxin, and cross-contamination from shared lab equipment. Structurally similar impurities are the hardest category to catch because HPLC-UV alone often can't resolve them from the target peptide.
Can HPLC-UV Alone Confirm Peptide Purity?
No. HPLC-UV separates by hydrophobicity, so epimers, deletion sequences, and other structurally similar impurities can co-elute with the target peptide and go undetected. Orthogonal methods like LC-MS, NMR, or SEC are needed to confirm identity and resolve impurities near identification thresholds.
Why Do Endotoxin Tests Need Matrix-Specific Validation?
A peptide can inhibit or enhance the LAL reaction used in gel-clot, turbidimetric, or chromogenic endotoxin assays, distorting the result. FDA and USP <85> guidance requires inhibition and enhancement controls run in the actual peptide matrix before any result can be trusted.
How Does USAPeptide Help Verify Peptide Quality?
USAPeptide's COA grading tool checks whether a supplied Certificate of Analysis actually supports its stated claims, and its molecular profile database helps researchers select the right orthogonal test for a given sequence. Sourcing referrals point toward ISO 17025-accredited testing as a baseline for lot verification.
What Should a Lab Do Immediately Upon Receiving a Peptide Shipment?
Inspect packaging condition, reconcile the physical lot against the COA, log receipt time and temperature history, and assign a unique accession number before the first aliquot is drawn. Quarantine any shipment with damaged packaging or a confirmed temperature excursion pending disposition review.