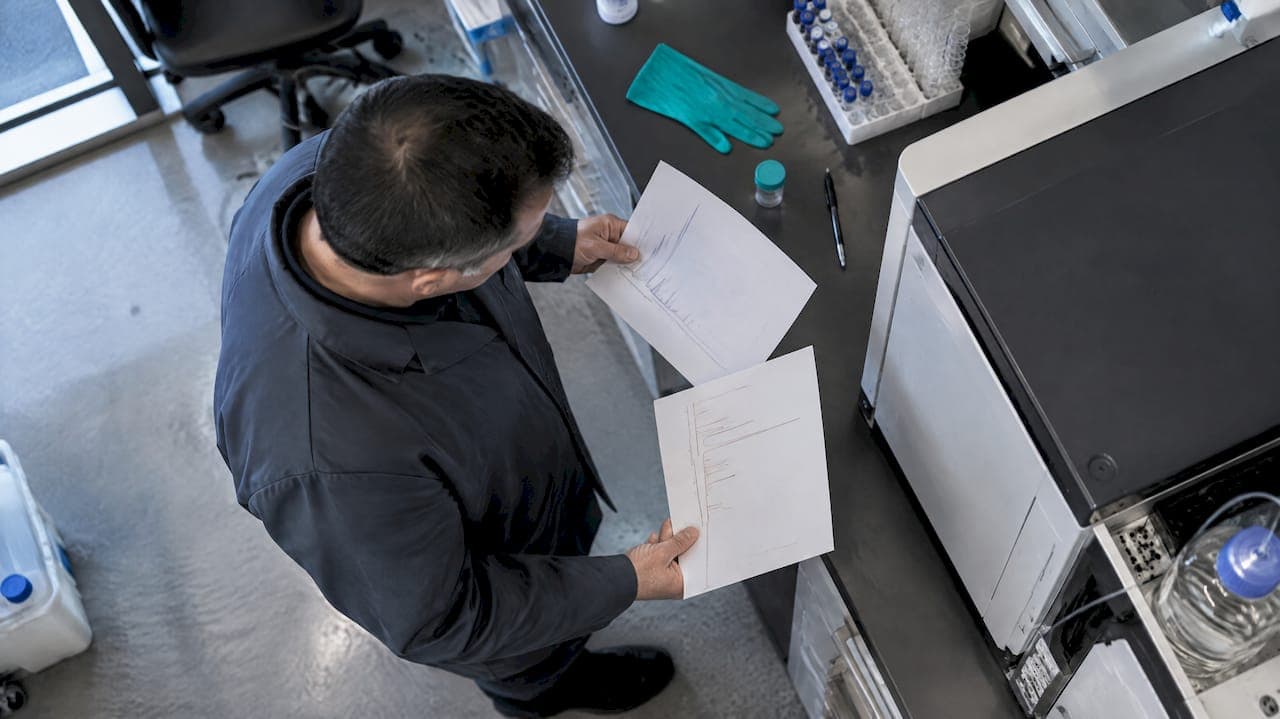
Batch-to-batch variability in peptide reagents refers to measurable differences in identity, purity, impurity content, counter-ions, residual solvents, or aggregation between production lots. The single most useful check on receipt of a new lot is to pull the Certificate of Analysis and confirm it states full test method parameters, a measurement uncertainty value, and at least one orthogonal identity test, which readers can cross-check using COA grading tools built for this purpose.
TL;DR:
- Confirm that peptide lot Certificates of Analysis include full test parameters, measurement uncertainty, and orthogonal identity verification methods before use.
- Be aware that small discrepancies in purity or impurity profiles may not be apparent from a single number and require multiple analytical techniques for confirmation.
- When comparing two lots, overlap in measurement uncertainty ranges often indicates equivalence, even if the purity percentages differ slightly.
- Assess laboratory accreditation scope carefully, ensuring the specific assay used for the lot is within its certified testing scope.
- Tighter procurement criteria, such as impurity thresholds and retention sample testing, reduce the risk of batch variability affecting experimental reproducibility.
- ✓Detailed molecular profiles
- ✓Peer-reviewed research summaries
- ✓COA grading tool
- ✓Verified Certificates of Analysis
Table of Contents
- How batch variability shows up on COAs and in the lab
- Analytical methods that reliably detect lot-to-lot differences
- Reading a COA: key fields, ISO/IEC 17025 scope, and measurement uncertainty
- Procurement and lab controls to limit variability and respond to it
- Weighing cost against risk when a lot looks inconsistent
- How USAPeptide.info supports COA verification and vetting
- FAQ
- Sources
- Authoritative primary documents and guidance
How batch variability shows up on COAs and in the lab
Variability rarely announces itself. It shows up as small inconsistencies that accumulate into a documentation gap or an experimental anomaly. A few COA-level red flags are worth scanning for before a lot ever reaches the bench:
- Missing method parameters, such as column type, mobile phase, or gradient, that make the stated purity percentage impossible to verify.
- No measurement uncertainty value attached to the purity or impurity result.
- A large purity shift between lots with no impurity identification explaining the difference.
- No mention of retention samples or prior lot history for comparison.
In the lab itself, signs tend to surface downstream: a bioassay that suddenly reads lower potency, a new or growing peak on an HPLC trace that was not present in the prior lot, a change in solubility or visible aggregation, or a mass balance that no longer adds up once water, counter-ion, and solvent content are accounted for. A single HPLC purity percentage, on its own, can mask exactly this kind of risk.
Analytical methods that reliably detect lot-to-lot differences
Relying on one number from one method is the most common mistake in comparative peptide evaluation. USP reference standard guidance recommends combining liquid chromatography-tandem mass spectrometry for identity with HPLC for purity and mass balance, because a single purity percentage is not enough to establish that two lots are consistent.
A practical sequence for verifying or rejecting a lot:
- Request the full HPLC related-substance method, including column chemistry, particle size, mobile phase composition, gradient, flow rate, and detection wavelength.
- Confirm identity through mass spectrometry or LC-MS/MS and, where available, peptide mapping against a characterized reference standard.
- Apply an orthogonal method, such as high-resolution MS, NMR, or ion chromatography, to resolve impurities that co-elute under the primary HPLC conditions.
- Check for aggregation or oligomer formation when the peptide's use case involves higher concentrations or extended storage.
- Ask for release, end-of-shelf-life, and, where relevant, stressed samples rather than a single release-only data point.
HPLC chromatographic parameters materially change peak shape and resolution. Unless two COAs report identical validated method parameters, their purity percentages are not strictly comparable, a point USP's peptide standards guidance makes explicit when describing how peptide quality attributes differ from small-molecule specifications.
Pro Tip: Ask for the raw chromatogram alongside the summary purity number. A clean-looking percentage can hide a shoulder peak that a summary table rounds away.

Reading a COA: key fields, ISO/IEC 17025 scope, and measurement uncertainty
A usable COA gives us more than a final number. It states the full method parameters, reports the result as a value plus an expanded measurement uncertainty, documents a decision rule for pass or fail, and names the identity methods used alongside any retention sample reference.
ISO/IEC 17025 accreditation does not mean every test a lab runs is accredited. ISO/IEC 17025:2017 requires accredited labs to define and apply decision rules for statements of conformity and to report measurement uncertainty where it affects interpretation. Accreditation should be read narrowly: a purchaser needs to confirm that the specific assay used for a given lot, such as the particular HPLC-UV method, actually falls within that lab's accredited scope rather than assuming blanket coverage.
When comparing two lot results, the uncertainty value matters as much as the result itself:
- A 98.5% purity result and a 97.9% purity result may not reflect a real difference if both carry an uncertainty wide enough to overlap.
- A documented decision rule tells us how a lab converts a measurement plus uncertainty into pass or fail, removing guesswork from the comparison.
- Comparing lots without accounting for uncertainty risks both false rejections of acceptable material and missed failures that should have been caught.
A deeper walkthrough of COA fields and how to apply them is available in our lab checklist for reading a COA.
Procurement and lab controls to limit variability and respond to it
Specifications that stop at a purity percentage leave the door open to exactly the variability described above. Tighter procurement language closes most of that gap.
- Set acceptance criteria beyond purity percent: impurity identification thresholds, a rule against new impurities above 0.10% without documented justification, aggregate limits, and counter-ion or residual solvent limits.
- Maintain retention samples and test new lots against a retained reference lot under identical method parameters rather than against a historical number alone.
- Require vendors to supply full method parameters and measurement uncertainty on every COA, and to disclose any change in manufacturing process or site between lots.
- Commission an independent ISO-accredited comparative study when a lot shows a borderline result, an unexplained impurity, or when the intended use carries higher reproducibility stakes.
FDA guidance on analytical procedures points to retention samples as one of the more time-saving tools in a comparative investigation, since they let a lab run a new analytical method against legacy material instead of depending solely on a vendor's prior statement. Regulators generally treat impurities at or above identified low levels as warranting identification, and new impurities or aggregates appearing above that level commonly trigger a root-cause review rather than a quiet acceptance.
Pro Tip: Build a one-page acceptance template before the next order arrives. Reviewing criteria after a questionable lot is already on the bench is the slowest way to resolve it.
Residual solvents deserve particular attention in this process, since they factor directly into mass balance; our guide to residual solvent testing covers the limits and reporting conventions involved.
Weighing cost against risk when a lot looks inconsistent
Full comparative testing is expensive, and not every borderline COA justifies it. A documented COA review, cross-checked against our COA grading tool and the lab's stated ISO-accredited scope, is usually enough to decide whether a result is a genuine concern or a reporting gap. Our recommended escalation ladder runs from document review, to targeted orthogonal testing on the specific peptide in question, to an independent ISO-accredited comparative study only when the first two steps leave the question unresolved. Keeping clear lot history and retention samples on file shortens every step of that process considerably.
— USAPeptide Team
How USAPeptide.info supports COA verification and vetting
Verifying a COA is faster with the right reference material on hand rather than starting from a blank page every time a new lot arrives, such as using an APYTFGQGTK peptide control. We built our COA grading tool to let labs pre-check a Certificate of Analysis against the fields covered above, and our peptide reference and glossary defines the technical terms that show up in method parameters and impurity reporting.
- Pre-check a COA before an order ships using our grading tool.
- Pull definitions for terms like mass balance or relative retention time from our glossary while reading a report.
- Export a COA summary for procurement records once a lot clears review.
When it is time to source compounds for a specific research area, our peptide reference pages list supplier referrals alongside the molecular data behind each compound.
This article is general information, not a substitute for advice from a qualified doctor. Consult a qualified healthcare professional about your own circumstances before acting on anything here.
FAQ
What counts as batch-to-batch variability in research peptides?
It refers to measurable differences in identity, purity, impurity content, counter-ions, residual solvents, or aggregation between production lots of the same peptide. These differences can affect experimental reproducibility even when each lot individually meets a purity specification.
Why isn't a high HPLC purity percentage enough to confirm consistency?
A purity percentage alone does not identify what the remaining impurity consists of, and USP reference standard guidance recommends pairing HPLC purity data with mass spectrometry-based identity testing. Two lots at the same stated purity can carry different impurity profiles that only an orthogonal method would reveal.
What does ISO/IEC 17025 accreditation actually guarantee?
ISO/IEC 17025 accreditation confirms a lab follows defined technical and management requirements for specific tests within its accredited scope, not for every test it performs. Buyers need to confirm the exact method used for their lot, such as a particular HPLC assay, is itself within that scope.
How should researchers compare two lot results that look different?
Compare the results alongside their measurement uncertainty rather than as bare numbers, since overlapping uncertainty ranges can mean there is no real difference between lots. A documented decision rule from the testing lab clarifies how a result plus uncertainty translates into a pass or fail outcome.
When is independent comparative testing worth the cost?
Routine COA review combined with retained reference samples resolves most other cases.
Sources
- USP: Peptide standards and reference materials
- Satra summary of ISO/IEC 17025:2017
- USP reference standards to support peptide quality
- FDA: Common deficiencies in comparative peptide impurity profile studies (guidance excerpt)
- FDA: Analytical procedures and methods validation for drugs and biologics
Authoritative primary documents and guidance
- FDA: Common deficiencies in comparative peptide impurity profile studies: impurity identification thresholds and comparative testing expectations.
- FDA: Analytical procedures and methods validation: method robustness and retention sample guidance.
- USP: Peptide standards and reference materials: peptide-specific quality attribute standards.
- USP reference standards for peptide quality: identity and purity testing recommendations.
- ISO/IEC 17025:2017 summary: decision rules and measurement uncertainty requirements.